
US FDA Medical Device Consultant &
510(k) Premarket Notification
As a premier medical device consultancy, we provide end-to-end USFDA (US FDA) market access support — from 510(k) and 510 k substantial equivalence strategy and predicate analysis to testing coordination, eSTAR dossier preparation, FDA submission, and deficiency response.
Regulatory Consulting
& IVD Expertise
Local Understanding
Our US FDA Submission Approach
Assess
Evaluate device, intended use, classification, applicable pathway & documentation gaps
Strategize
Predicate strategy, testing strategy & regulatory roadmap
Test
Test planning & laboratory coordination
Prepare
eSTAR dossier & supporting technical documentation
Submit
FDA submission through CDRH Portal
Respond
Address FDA questions, deficiencies & additional information requests
Maintain
Post-clearance support, change assessment & lifecycle management
Specialized US FDA Regulatory Solutions
End-to-end support across every phase of device classification, testing, clearance, and post-market compliance.
Product Classification & Regulatory Pathway Assessment
Before testing begins or a dossier is drafted, we establish the device's regulatory identity: classification, regulation number, product code, applicable controls, exemption status and likely premarket pathway. This early assessment prevents costly work from being built around the wrong FDA route.
Learn more510(k) Premarket Notification Support
We structure the 510(k) around the substantial equivalence decision FDA must make. Classification, predicate eligibility, technological differences, evidence requirements and labeling are aligned before the eSTAR submission is built.
Learn morePredicate & Substantial Equivalence Strategy
We evaluate whether a proposed predicate can withstand FDA's substantial equivalence analysis, then trace every meaningful technological difference into the evidence needed to support it.
Learn moreTesting Strategy & Laboratory Coordination
We design the testing plan backward from the FDA decision the evidence must support — challenging configuration, worst-case rationale, standards, acceptance criteria and whether the resulting data will actually answer the regulatory question.
Learn moreeSTAR Dossier Preparation
We treat eSTAR as the container for the regulatory argument — not as the strategy itself. The dossier is built only after pathway, predicate and evidence requirements are clear, then checked for cross-section consistency.
Learn moreFDA Q-Submission & Pre-Submission
We determine which uncertainties are worth taking to FDA, frame focused questions and provide enough technical context for FDA feedback to be actionable before major testing or submission decisions are locked.
Learn moreDe Novo & PMA Support
For novel or higher-risk devices, we build the regulatory strategy around the evidence FDA will need to reach a classification or approval decision, integrating risk, controls, non-clinical evidence, clinical evidence and FDA interaction.
Learn moreFDA Additional Information & Deficiency Response
We deconstruct each FDA question to identify the underlying regulatory concern, trace it back through the submission and determine whether the real issue is evidence, study design, predicate logic, labeling, consistency or presentation.
Learn moreQMSR & FDA Inspection Preparation
We test whether the quality system can produce objective evidence when an FDA investigator follows a real transaction through the organization — not merely whether the required SOPs exist.
Learn moreUS Agent Services
We provide US Agent representation as part of a broader regulatory interface — supporting FDA communication, registration coordination and escalation when the foreign manufacturer needs a reliable US regulatory contact.
Learn moreFDA Establishment Registration & Device Listing
We support initial registration, device listing and annual maintenance while checking that the administrative registration activity remains aligned with the device's actual regulatory status and market authorization pathway.
Learn morePost-Clearance & Device Lifecycle Support
We assess the regulatory impact of device changes before implementation — focusing on whether the modification affects intended use, technological characteristics, safety, effectiveness, performance or the basis of the original clearance.
Learn morePlanning for Global Market Access
We plan regulatory evidence across target markets so common testing and technical documentation can be leveraged where scientifically and regulatorily appropriate, while market-specific gaps are identified early.
Learn moreCurrent FDA reference
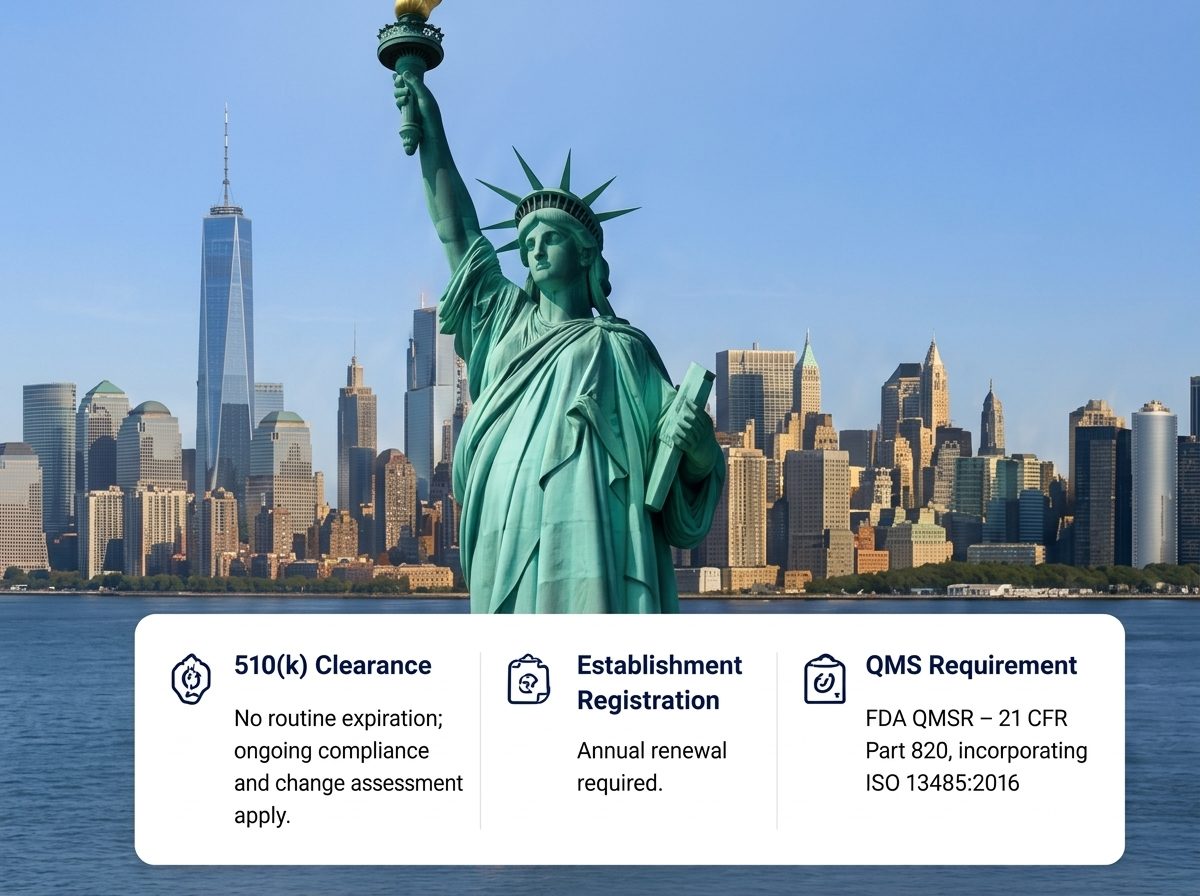
FDA’s Quality Management System Regulation became effective on 2 February 2026 and incorporates ISO 13485:2016 by reference. The applicable premarket and postmarket requirements still depend on the device, establishment, and pathway.
Why Manufacturers Choose
NKB Regovanta
- Boutique regulatory consulting with senior level, hands-on support
- Strategy before submission – not just dossier preparation
- Medical Device & IVD expertise across all risk classes
- End-to-end FDA support from strategy to clearance and beyond
- Testing coordination with trusted laboratory network
- Global perspective to support multi-market regulatory strategies
Ready to Enter the US Market?
Let our US FDA regulatory experts guide you through the right pathway with confidence.
Talk to a Regulatory Expert2018
Experience Since
Regulatory & Quality Experience
350+
Completed
Regulatory & Quality Projects
21+
Supported
Global Clients
15+
Supported
International Countries
4
Focus Areas
Core Industries
Global
India Based
Connected Network of Experts